
The limiting factor in reading the genetics of a diseased organ is not sequencing depth; it is knowing which of a cell's two copies of a gene is actually being used. Each of us carries two versions of every gene on the non-sex chromosomes, and the textbook assumption is that both contribute roughly equally. Often they do not. When one copy dominates, the phenomenon is called allele-specific expression, and it captures regulation happening within a single cell rather than averaged across a tissue. That makes it a sensitive instrument, and a treacherous one.
A team at Boston Children's Hospital has now mapped that imbalance genome-wide in kidney biopsies from patients with proteinuric disease, in which the kidney's filters leak protein into the urine. Two findings stand out. Diseased kidneys showed considerably more allelic imbalance in the glomeruli, the filtering units themselves, than in the surrounding tubules, a split absent in control kidneys. And the patients with the widest split did measurably better: they were 65% less likely to reach kidney failure or lose 40% of their filtration capacity.
Why it matters: Proteinuric disorders account for up to 90% of end-stage kidney disease worldwide, and clinicians still have poor tools for telling early which patients will deteriorate. A marker separating the resilient from the vulnerable from a single biopsy would change how they are monitored.
The study, by Onuchic-Whitford et al. at Boston Children's Hospital, with collaborators at Brigham and Women's Hospital, Harvard Medical School, the Broad Institute and the University of Michigan, appears in Science Advances. It draws on the Nephrotic Syndrome Study Network, or NEPTUNE, unusual in pairing whole-genome sequencing with RNA sequencing from biopsy tissue microdissected into its two compartments before being read.
Why the measurement is harder than it sounds
Much of the paper is about not fooling yourself. Allelic imbalance is inferred by counting reads carrying one variant against those carrying the other, and nearly every step of that pipeline can manufacture a false signal. Genotyping errors make a site look heterozygous when it is not. Reads from genes that physically overlap on the chromosome get credited to both, a problem affecting roughly 48% of expressed genes here. The authors built filters for each: blacklisting variants that behaved impossibly, discarding 18 samples whose DNA and RNA did not match, and writing a new tool to reassign reads in overlapping regions, which resolved 75% of genes cleanly. Before filtering, an average of 302 genes per sample would have appeared imbalanced purely from bad genotypes.
The same rigour supports the paper's other broad claim. Imbalance was overwhelmingly private, with most events appearing in only a handful of individuals, but the common ones were not arbitrary. Among frequently shared imbalanced genes in the glomeruli, 86.2% traced to a known sequence-based cause: expression or splicing quantitative trait loci, or genomic imprinting. Where the mechanism was unknown, those genes were specifically enriched for rare variants in their promoter regions, the kind population-scale studies lack the numbers to detect. Individual-specific imbalance, long written off as stochastic, looks instead like each person's own regulatory architecture.
The compartment split, and what it predicted
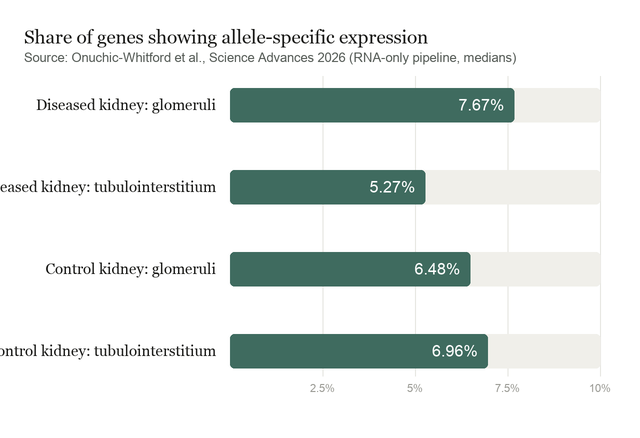
In diseased kidneys the median share of genes showing allelic imbalance was 11.3% in glomeruli against 7.0% in tubulointerstitium, with 177 of 206 patients showing the glomerular excess. In 31 control kidneys the two compartments were indistinguishable, at 6.48% versus 6.96% (p = 0.68). The authors reduced this to one number, the glomerular-to-tubular ratio, and split 194 patients at its midpoint.
The high-ratio group progressed more slowly, with an adjusted hazard ratio of 0.35. They also reached complete remission of their protein leakage sooner (hazard ratio 1.58, p = 0.023), and six of the seven patients who went on to kidney failure were in the low-ratio group. Scarring measures pointed the same way without reaching significance: global glomerulosclerosis at p = 0.083 and interstitial fibrosis at p = 0.051. Two controls make the result worth attention. Neither compartment's imbalance predicted anything on its own; only the ratio did. And deconvolution found no difference in cell-type composition between groups, so the high-ratio kidneys were not simply the ones with more surviving podocytes.
Gene expression suggested why. Comparing the groups turned up 181 differentially expressed genes in glomeruli and none in tubules. The high-ratio group had ribosome biogenesis and oxidative phosphorylation turned up, immune signalling and extracellular-matrix pathways turned down; 10 of the 66 downregulated genes promote kidney fibrosis. That is the profile of a cell working hard, not one losing control.
What the study can't say yet
The ratio is a correlation found once, in one cohort, with no independent replication, and the authors are explicit that they could not attempt it because comparably paired datasets barely exist. Validating the metric, and establishing how sensitive it is to sample handling and sequencing depth, stands between this and any clinical use.
The control comparison carries an asymmetry worth naming. Those kidneys had RNA sequencing but no paired genomes, so they were analysed with a weaker RNA-only pipeline that systematically misses the most extreme imbalance. The patient samples were re-run the same way for a fair comparison and the split held, but the cohorts also differ in age, ancestry and read depth.
Bulk sequencing of a microdissected compartment is also not single-cell sequencing. The authors note that cell-state diversity, localised clonal expansion and transcriptional bursting could each generate apparent imbalance, and cannot be ruled out without single-cell measurement. On causation they are deliberately restrained, concluding that the observed increase in ASE is a by-product of adaptive transcriptional and metabolic activity, which may confer glomerular resilience to proteinuric injury, the authors write. A by-product, not a cause.
Quick questions
Is this a test a patient could get? Not yet. The team did derive a simpler score from ordinary expression levels, bypassing the haplotype counting; it separated the groups reasonably well (area under the curve 0.77) but only trended toward predicting kidney survival (p = 0.08).
Why would unequal gene copies ever be a good sign? They are not good in themselves. The reading here is that a cell mounting a vigorous transcriptional response expresses more genes more strongly, and stronger expression makes existing regulatory differences between the copies easier to detect.
What is the one-line takeaway? In diseased kidneys, more lopsided gene expression in the filtering units tracked with better outcomes, apparently as a side effect of an active protective response rather than as its cause.
Onuchic-Whitford et al. "The landscape of allele-specific expression in human kidneys." Science Advances, 2026. doi.org/10.1126/sciadv.aeg3462
PubMed PMID: 42826188.
Image: renal corpuscle, light micrograph. Ed Uthman, CC BY-SA 2.0, via Wikimedia Commons.
Comments